RoseTTAFold: Three-Track Biomolecular Modeling
Minkyung Baek, Frank DiMaio, David Baker et al. (Baker Lab, 2021)
Baek, M., DiMaio, F., Anishchenko, I., ... & Baker, D. (2021). Accurate prediction of protein structures and interactions using a three-track neural network. Science, 373(6557), 871-876.
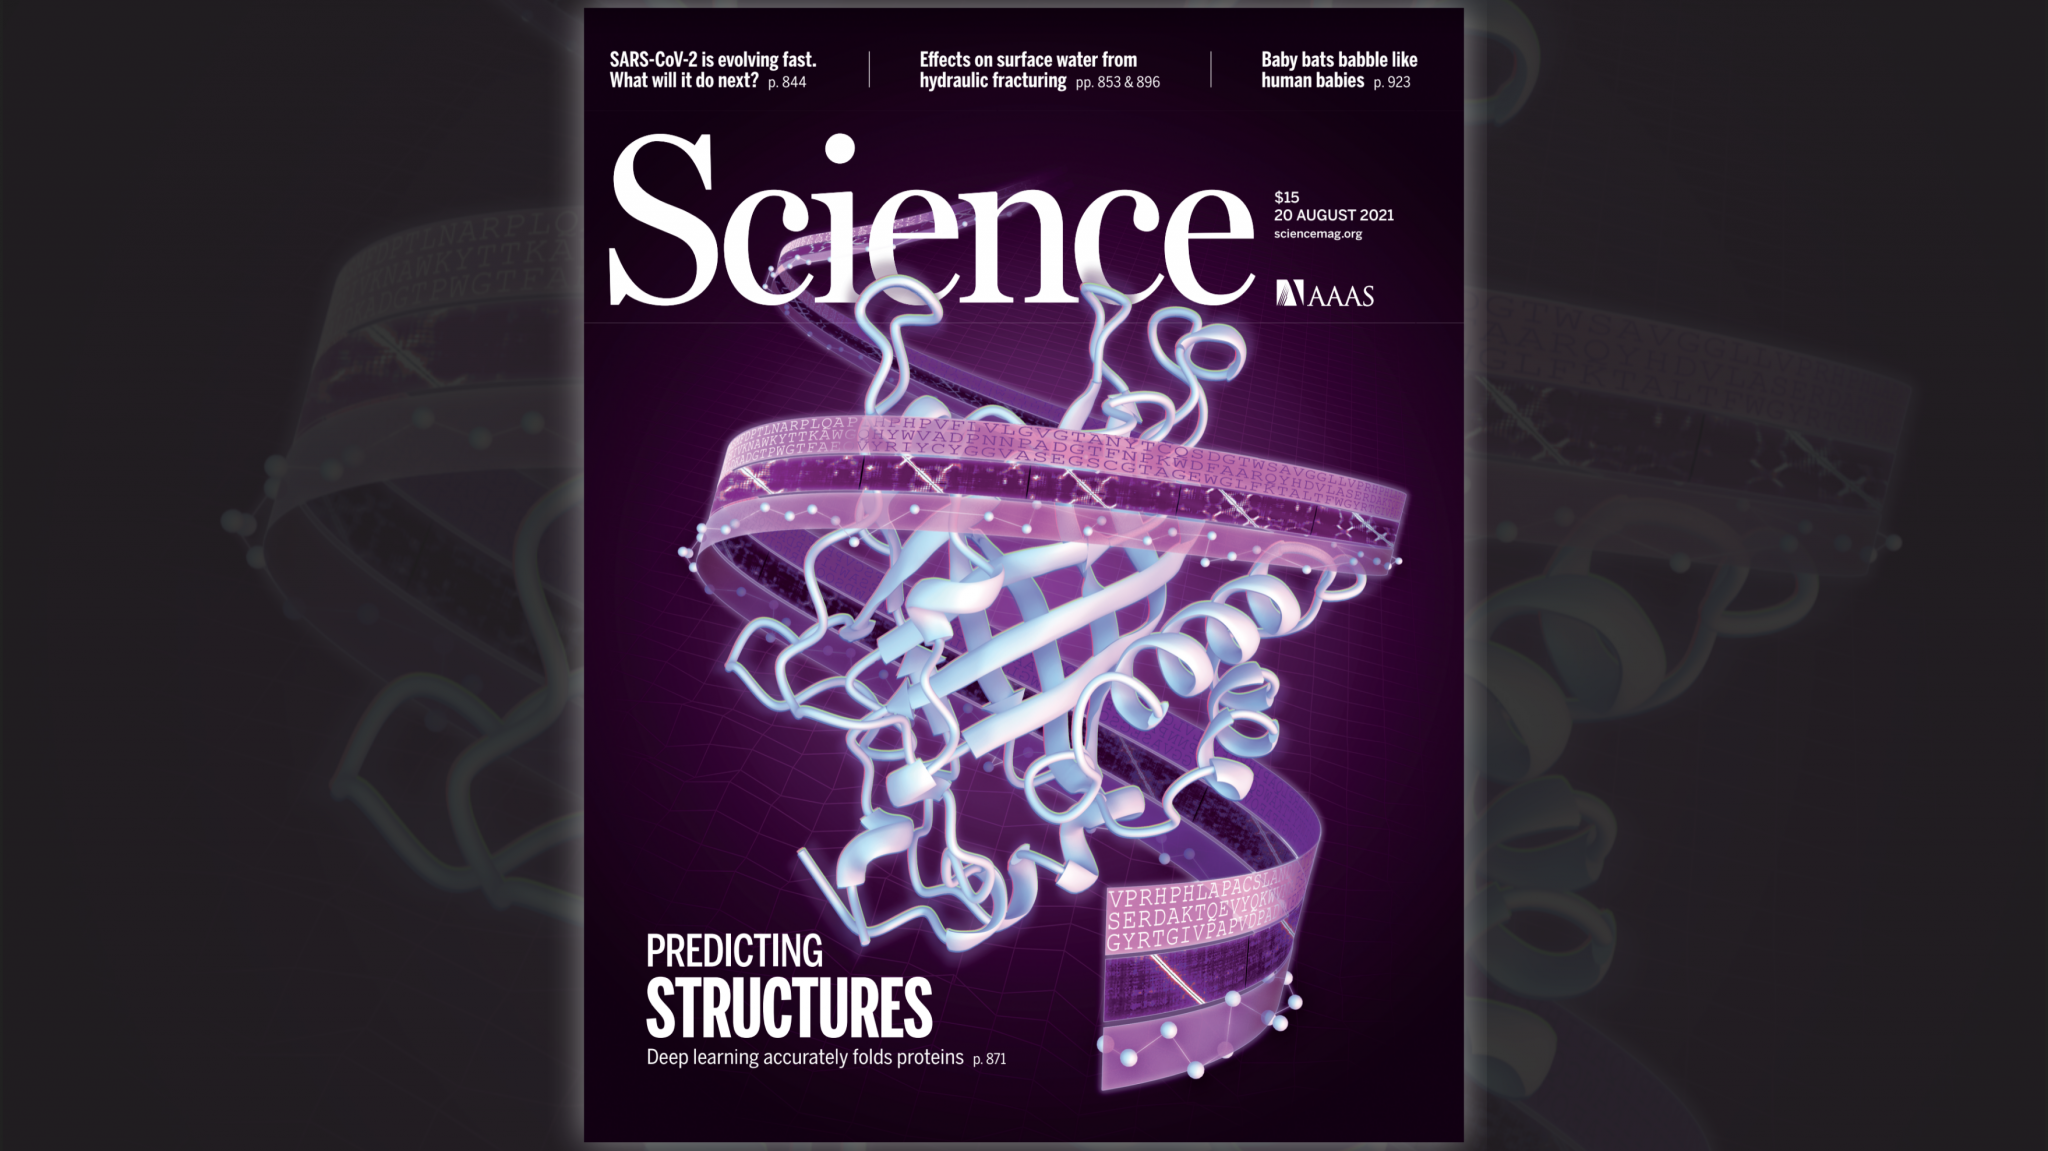
In 2021, researchers at the David Baker laboratory introduced RoseTTAFold, an open-source three-track neural network that achieved near-experimental accuracy in predicting 3D protein structures and macromolecular complexes. Prior to this work, computational structural biology operated through segmented stages: 2D neural networks predicted inter-residue distance distributions, followed by separate physical energy minimization packages (such as Rosetta) to build 3D coordinates. RoseTTAFold demonstrated that by coupling 1D sequence patterns, 2D distance maps, and 3D coordinate space into a single continuous feedback loop, neural networks can rapidly solve monomeric structures, multi-chain assemblies, and difficult crystallographic datasets.
The Gap in Decoupled Structural Pipelines
Before CASP14, protein modeling pipelines suffered from a disconnect between statistical contact prediction and physical 3D assembly. A neural network would process a multiple sequence alignment to generate pairwise distance histograms, and external gradient-based optimization tools would attempt to fold the chain into those constraints. Because information flowed strictly in one direction, emerging 3D spatial geometry could never correct mistaken co-evolution predictions in the sequence alignment. RoseTTAFold addressed this limitation by designing an architecture where sequence, pairwise contacts, and 3D Euclidean coordinates communicate bidirectionally at every layer.
The Three-Track Simultaneous Architecture
RoseTTAFold organizes representation learning across three parallel, interconnected tracks:
- 1D Sequence Track: Tracks multiple sequence alignment features and residue identities.
- 2D Pair Track: Tracks pairwise spatial relationships, inter-residue distance distributions, and relative orientations between all residue pairs.
- 3D Coordinate Track: Directly maintains and updates rigid backbone frames and atomic coordinates in 3D Cartesian space.
Rather than propagating information sequentially, information flows continuously between all three dimensions:
- 1D to 2D: The outer product mean of sequence features updates the pair matrix.
- 2D to 1D: Pair features apply attention biases across 1D sequence positions.
- 2D to 3D: Pair distances and orientations guide SE(3)-equivariant coordinate updates.
- 3D to 2D: Inter-residue distances calculated directly from current 3D coordinates are fed back to update the 2D pair representation.
The 3D track utilizes SE(3)-equivariant transformations, ensuring that when a molecular structure rotates or translates in space, internal coordinates rotate and translate identically, preserving physical consistency.
Complex Modeling and Experimental Crystallography
RoseTTAFold was validated across blind CASP14 targets and continuous CAMEO benchmarks, achieving average TM-scores above 0.80 and closely matching the performance of AlphaFold 2 while requiring significantly fewer computational resources. The system solved complex structures in minutes on a single workstation GPU.
Beyond single chains, RoseTTAFold excelled at predicting protein-protein interaction complexes, heterodimers, and multi-chain assemblies directly from paired alignments without requiring experimental docking templates. The predicted structures proved accurate enough to solve previously intractable X-ray crystallography molecular replacement problems and cryo-EM density maps.
Open-Source Foundations for Generative Biology
The release of RoseTTAFold democratized deep-learning structural biology by providing an accessible, modular codebase to the broader scientific community. By showing that 1D, 2D, and 3D biological features can be trained jointly with continuous feedback, RoseTTAFold provided the core architectural foundation for subsequent generative frameworks, including RFdiffusion. It demonstrated that modeling the physical world requires architectures that let geometric reality directly guide sequence interpretation.
Dive Deeper
Accurate Prediction of Protein Structures (Science)
Science • docs
Explore ResourceRoseTTAFold Official GitHub Repository
GitHub • code
Explore ResourceRobetta Structure Prediction Server
Baker Lab • docs
Explore Resource
Discussion
0Join the discussion
Sign in to share your thoughts and technical insights.
Loading insights...
Recommended Readings
The author of this article utilized generative AI (Google Gemini 3.1 Pro) to assist in part of the drafting and editing process.